FDA发布细胞与基因疗法最终指南:要求生产企业把效价保证贯穿产品全生命周期
美国食品药品监督管理局(FDA)11月2日发布针对细胞与基因疗法(CGT)生产企业的最终指南,要求企业实施一套全面、以科学和风险为基础的「效价保证策略」(Potency Assurance Strategy),在产品到达患者之前,检验并确认每一批次都达到既定的效价标准、能实现预期的治疗作用。
这份名为《细胞与基因疗法产品的效价保证》的最终指南取代了 FDA 2011 年的效价检测指南,适用于治疗癌症、罕见病等严重乃至危及生命疾病的疗法。按法律要求,生产企业须从产品获批时起、直至上市后监测阶段,持续验证产品按预期发挥作用;新指南给出了 FDA 当前的建议做法。
「接受细胞与基因疗法的患者往往在与严重或危及生命的疾病作斗争,他们应当确信给自己使用的产品会按预期发挥作用,」FDA 生物制品评价与研究中心主任 Karim Mikhail 说,「这份指南体现了 FDA 的承诺:确保美国始终是世界上最适合创新的地方,并让创新与最高的产品质量和患者安全标准相匹配。」
指南以灵活、基于风险的方式,建议企业把效价考量融入研发与商业化的每个阶段——从早期临床研究申请(IND)、生物制品许可申请(BLA)获批,到上市后的持续验证;内容涵盖原材料变异控制、生产过程检测,以及确认每批产品生物活性与预期疗效相符的实验室检测,并对保质期短的产品、病毒载体基因疗法与患者个体化疗法给出专门考量。FDA 将于12月10日举办网络研讨会,帮助相关方理解并落实这份指南。
(据美国食品药品监督管理局发布的新闻稿)
FDA Issues Final Guidance on Potency Assurance for Cell and Gene Therapies
The U.S. Food and Drug Administration (FDA) on November 2 issued final guidance for manufacturers of cellular and gene therapy (CGT) products, recommending a comprehensive, science- and risk-based Potency Assurance Strategy to test and verify — before a product reaches a patient — that every batch meets a defined standard of potency to do what the product is supposed to do therapeutically.
The final guidance, "Potency Assurance for Cellular and Gene Therapy Products," replaces the FDA's 2011 potency testing guidance for CGT products and applies to treatments for serious and often life-threatening conditions such as cancers and rare diseases. By law, manufacturers are expected to verify that their products work as intended, from the time of approval through post-approval monitoring.
"Patients who receive cellular and gene therapies are often battling serious or life-threatening conditions, such as cancers and rare diseases, and they deserve the assurance that the product administered to them will perform as intended," said Karim Mikhail, Director of the Center for Biologics Evaluation and Research. "This guidance reflects the FDA's commitment to ensuring America remains the best place in the world to innovate — and that innovation is matched by the highest standards of product quality and patient safety."
The guidance offers flexible, risk-based recommendations covering every stage from early-phase investigational new drug (IND) submissions through biologics license application (BLA) approval and continued post-market verification, including controls for variability in raw materials, testing during manufacturing, and lab tests confirming each batch's biological activity. It also addresses products with short shelf lives, viral vector gene therapies and patient-specific therapies. The FDA will host a webinar on December 10, 2026, to help stakeholders implement the guidance.
(Source: U.S. Food and Drug Administration news release)
来源:GSM·美国(原文链接)
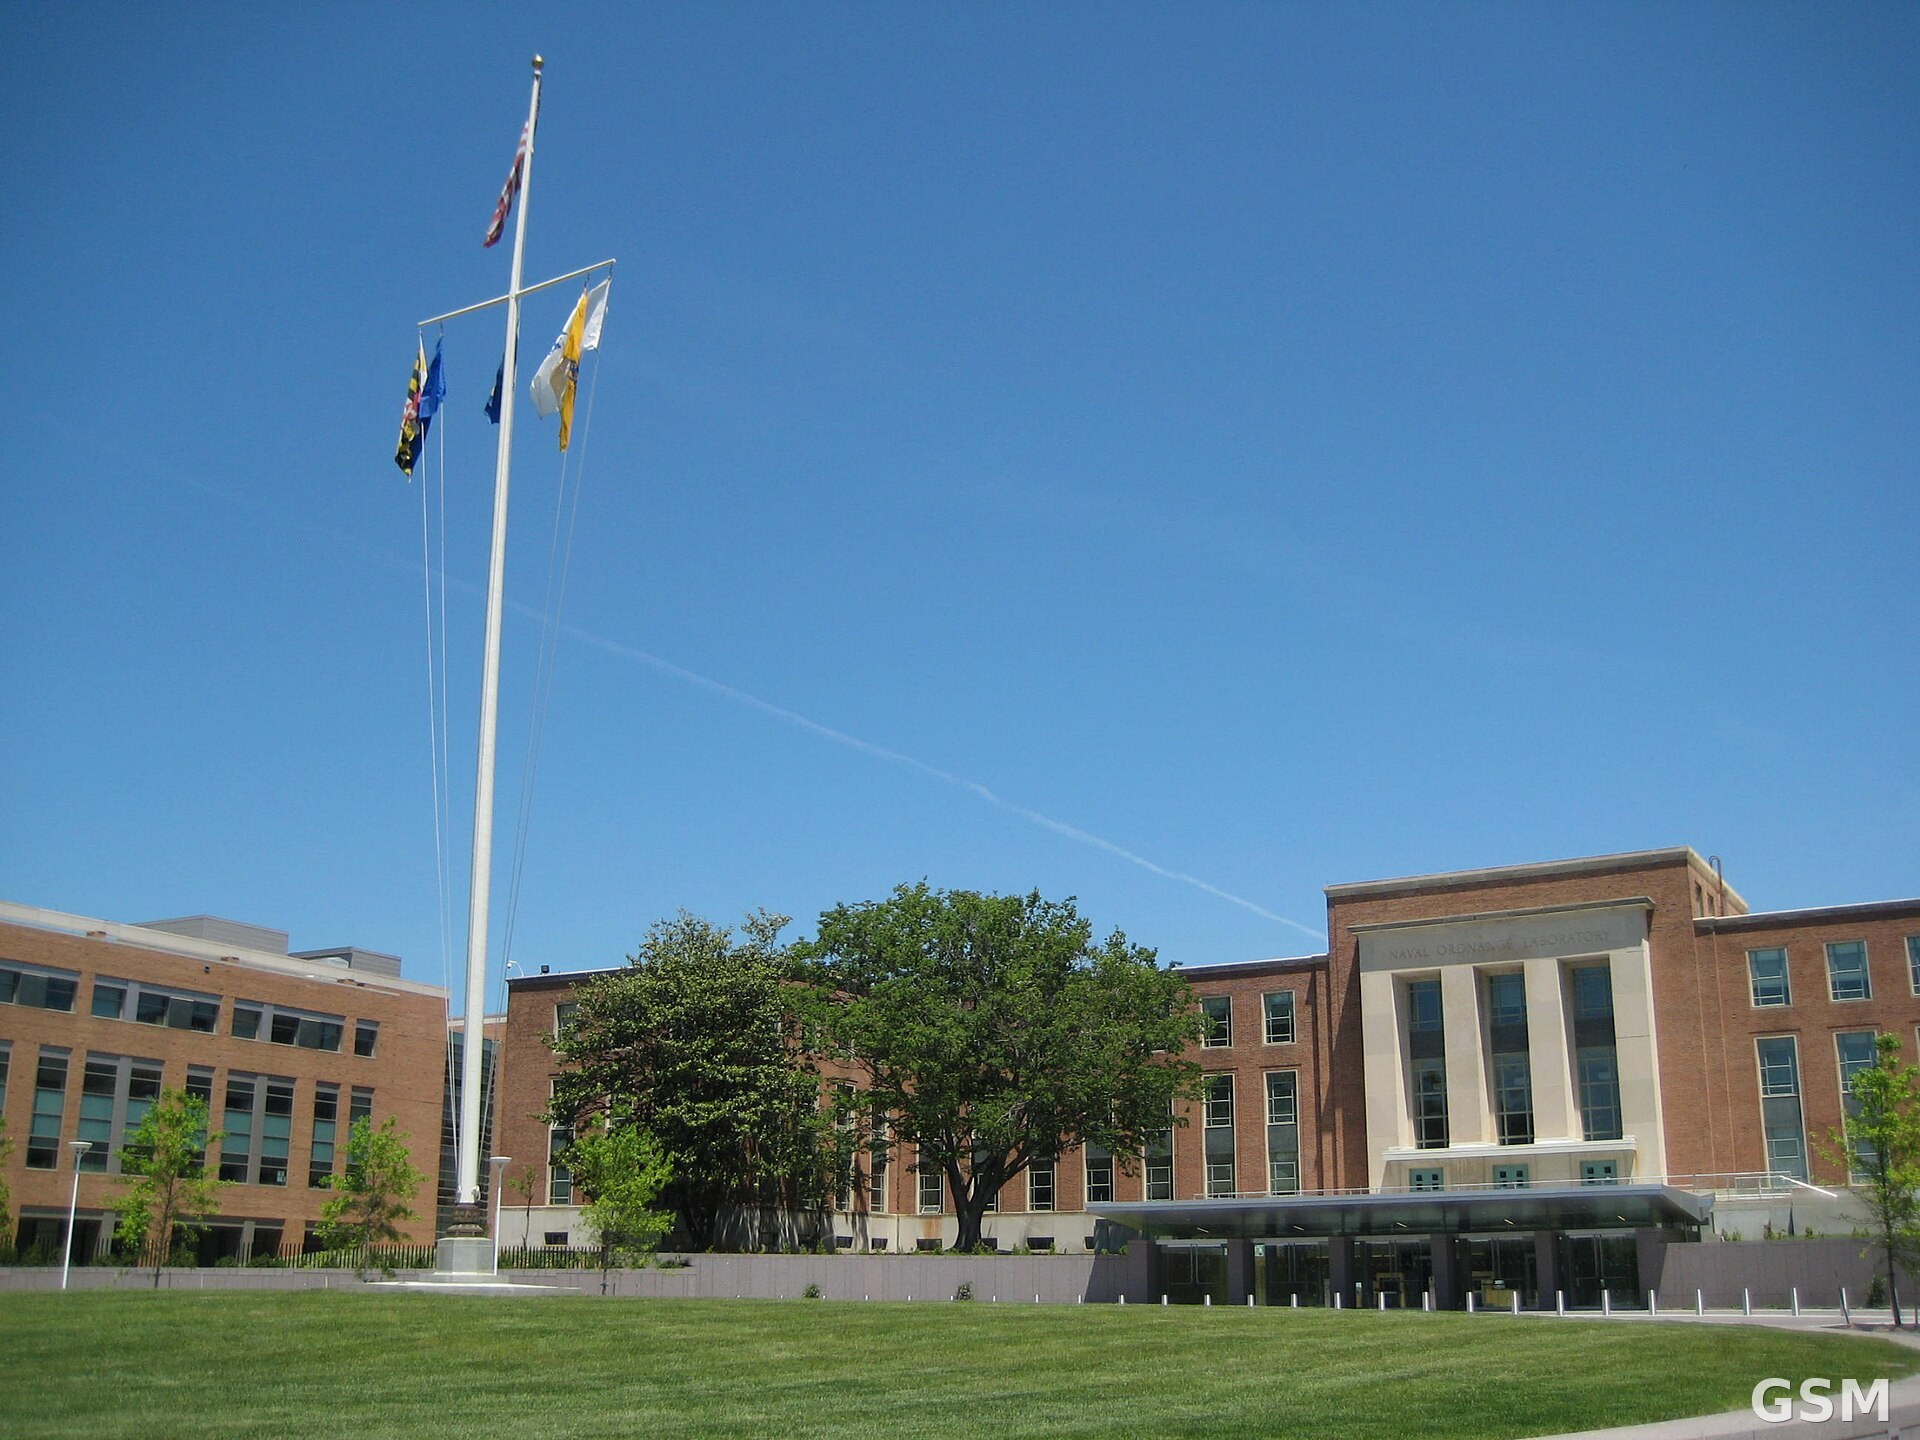
图片来源:图:美国食品药品监督管理局(FDA)位于马里兰州 Silver Spring 的 White Oak 园区总部大楼。FDA 官方图,公共领域,经 Wikimedia Commons。 / Photo: FDA headquarters buildings at the White Oak campus in Silver Spring, Maryland. U.S. Food and Drug Administration, public domain, via Wikimedia Commons.
GSM Global · 本篇英文版 English edition of this story