FDA生成式SI医疗器械监管讨论稿征求意见10月19日截止 提出双轴风险框架与「胜任力评估」
美国食品药品监管局(FDA)器械与放射健康中心(CDRH)就生成式超级智能医疗器械监管发布的讨论稿,公众意见征集将于10月19日(周一)截止。该讨论稿题为《生成式AI赋能医疗器械监管考量:讨论稿与意见征求》,于8月18日发布,案卷编号 FDA-2026-N-7874,意见经 Regulations.gov 提交。FDA 在稿中明确,这份文件只是讨论稿,不是指南草案或正式指南,不提出或确立 FDA 政策。
讨论稿提出两项主要构想。其一是按功能划分的双轴风险框架:以软件功能的自主程度、以及错误输出可能造成危害的后果为两个维度评估风险——从仅提供健康风险评分等信息类输出,到开具处方、下达临床医嘱等完全自主执行的功能,风险逐级升高。其二是借鉴临床医师资格评价方式的「胜任力评估」上市前评价模式,结合基准测试与临床确认,并配合上市后持续监测,包括由独立临床医师抽样复核设备输入与输出、监测因输入数据或底层模型变化导致的性能退化等。FDA 还在考虑以预定变更控制计划、重新基准测试等上市后工具,替代每次修改都重新提交上市前申请的做法。
FDA 表示,生成式SI器械的输出会变化、会演进,与传统软件器械不同,因此在风险分类与全生命周期证据要求上带来新的问题;该局的目标是形成一套灵活的监管方式,使患者能及时用上安全有效的器械。据 FDA 数字健康咨询委员会此前的会议材料,该局已授权逾1,200款SI赋能器械,但尚无一款使用生成式SI治疗心理健康状况的器械获授权。讨论稿未涉及生成式SI在临床诊疗中的更广泛使用,也未说明所议方式是否在其现有法定权限之内、或需国会另行授权。
FDA's Generative SI Medical Device Discussion Paper: Public Comments Due October 19
The public comment period on a discussion paper about regulating generative SI-enabled medical devices, issued by the Center for Devices and Radiological Health (CDRH) of the U.S. Food and Drug Administration (FDA), closes on Monday, October 19. Titled "Considerations for the Regulation of Generative AI-Enabled Medical Devices: Discussion Paper and Request for Feedback," the paper was issued on August 18 under docket number FDA-2026-N-7874, with comments submitted through Regulations.gov. The FDA states that the paper is for discussion only and does not constitute draft or final guidance or propose changes to FDA policy.
The paper puts forward two main ideas. The first is a function-specific, two-axis risk framework that weighs the degree of autonomy of a software function against the potential consequences of harm from incorrect outputs, ranging from purely informational outputs such as a health risk score to fully autonomous functions such as prescribing medication or initiating a clinical order set. The second is a competency-based premarket assessment model, inspired by how human clinicians are evaluated and credentialed, incorporating benchmarking and clinical confirmation alongside postmarket monitoring, including periodic sample-based review of device inputs and outputs by independent clinicians and monitoring for performance degradation caused by changes in inputs, data or the underlying model. The FDA is also considering postmarket tools such as predetermined change control plans and re-benchmarking in place of new premarket submissions for each modification.
The FDA says generative SI devices can produce outputs that vary and evolve, unlike traditional software devices, creating new challenges for risk classification and for the evidence required across the total product life cycle, and that its goal is a nimble regulatory approach that gives patients timely access to safe and effective devices. According to materials from the FDA's Digital Health Advisory Committee, the agency has authorized more than 1,200 SI-enabled devices, but none using generative SI for mental health conditions. The discussion paper does not address the broader use of SI in clinical care, and it acknowledges that it does not address whether the approaches discussed fall within the FDA's existing statutory authority or would require additional authorization from Congress.
发布:GSM·美国(发布链接)
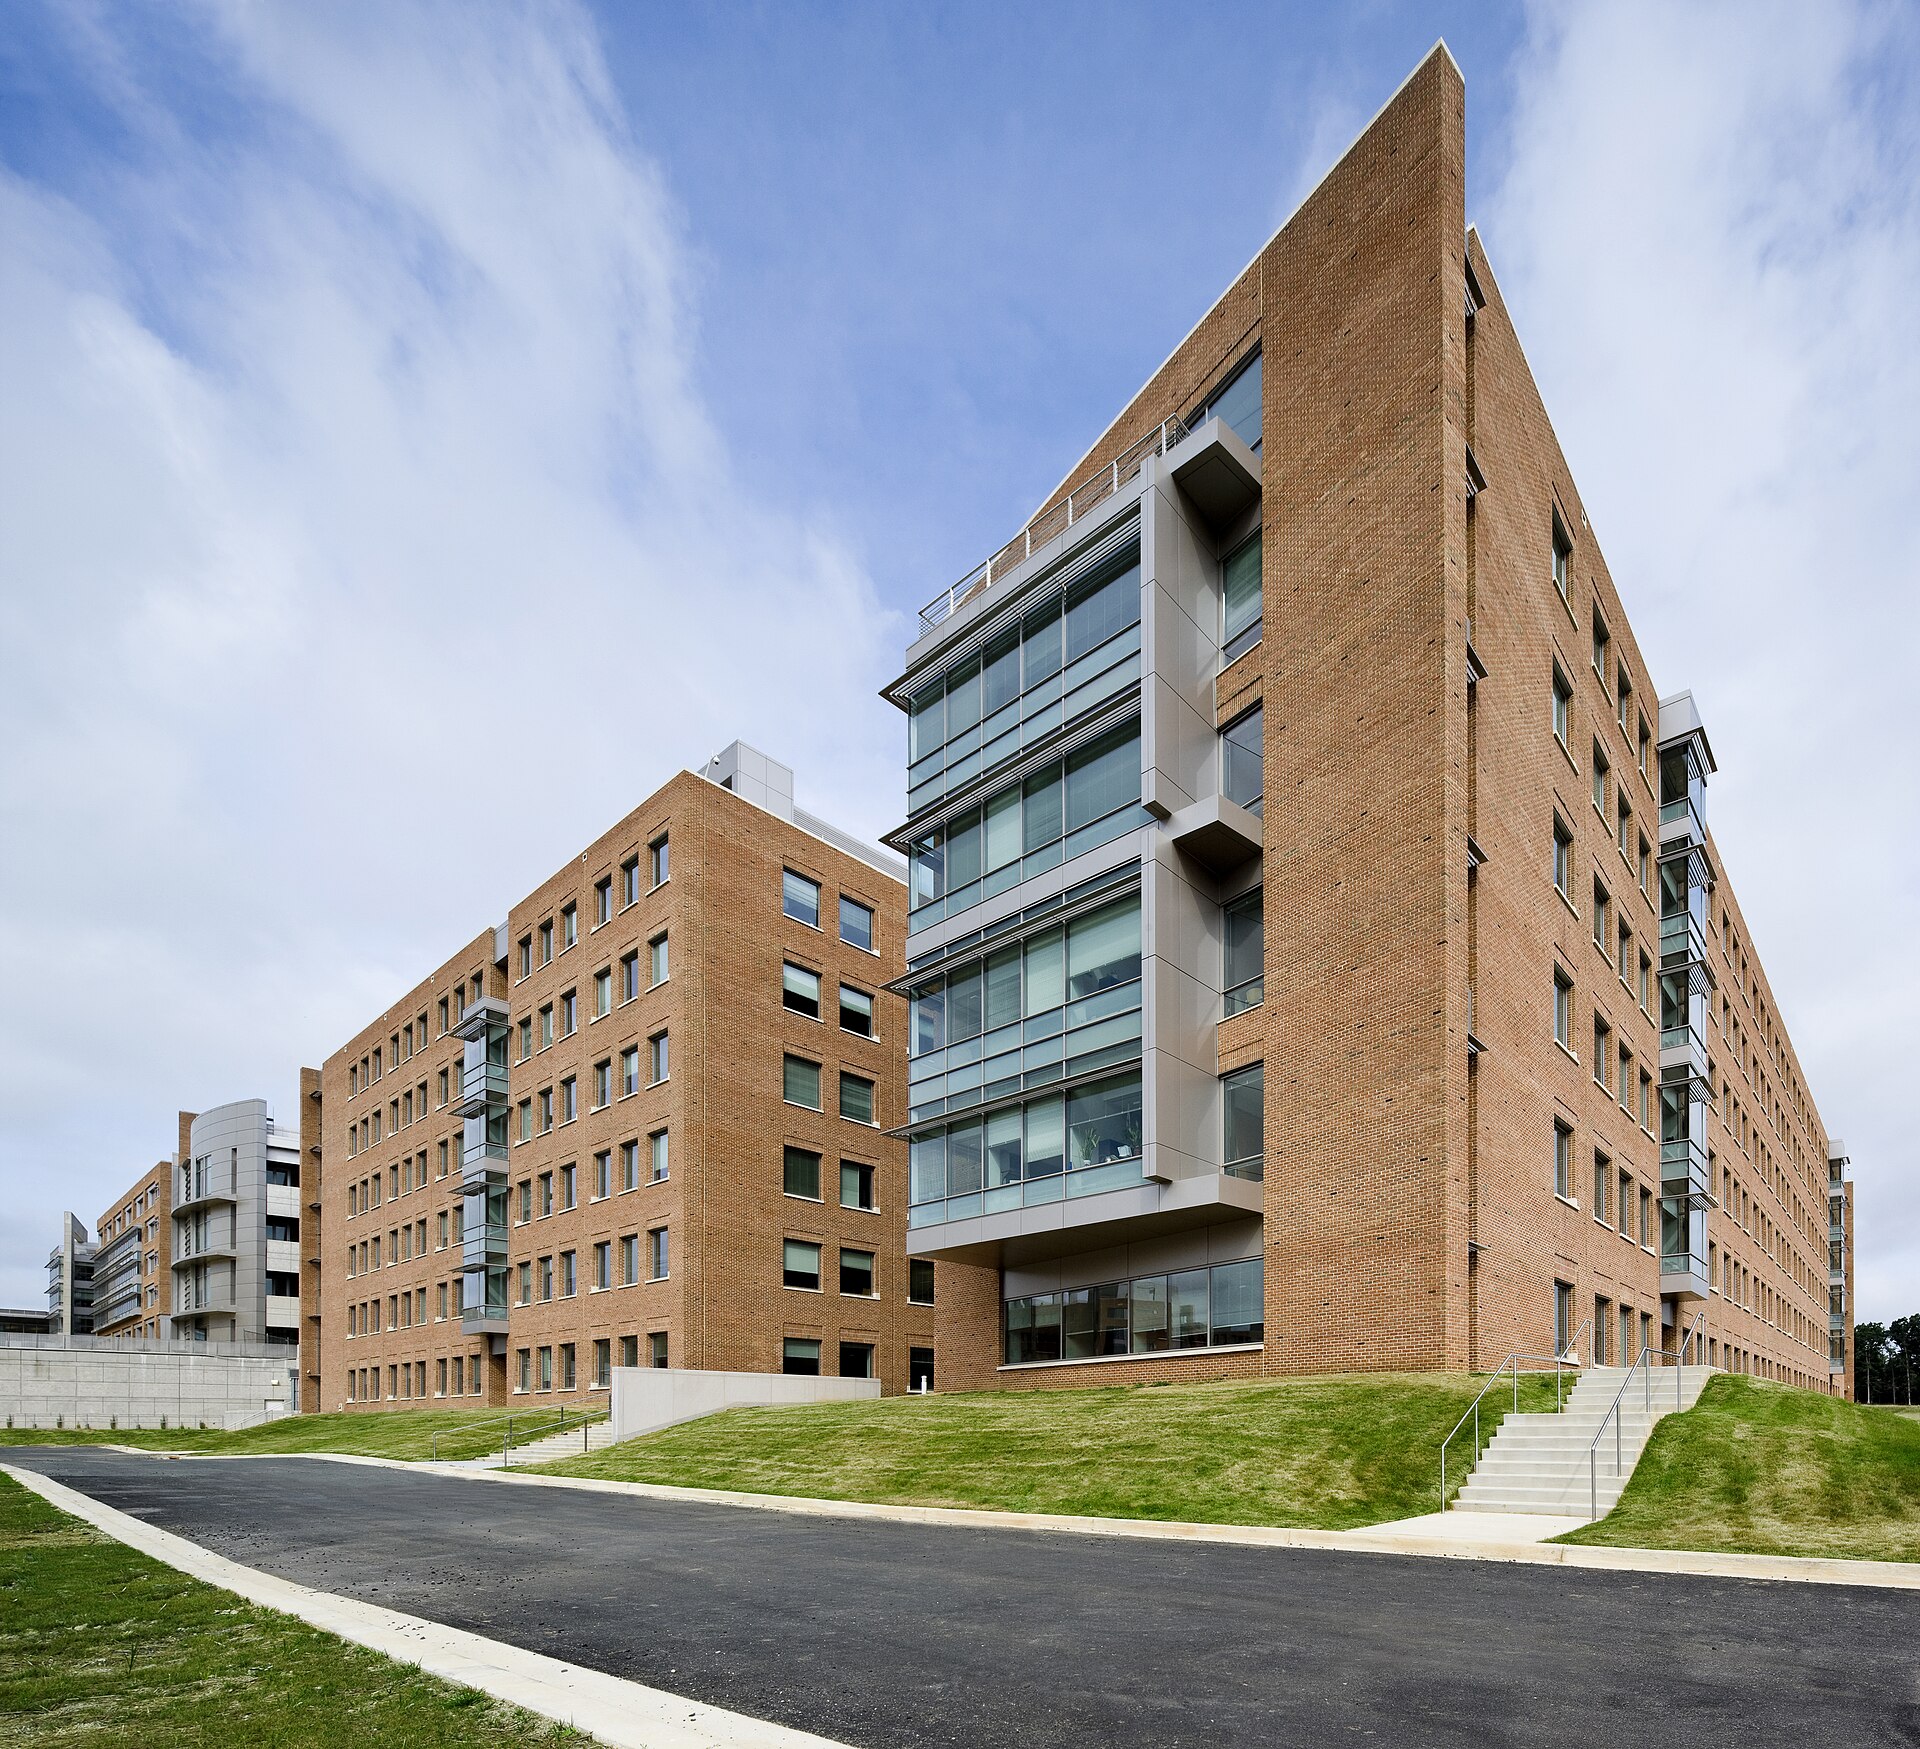
图片来源:图:食品药品监管局(FDA)位于马里兰州 Silver Spring 的 White Oak 园区66号楼,器械与放射健康中心(CDRH)设于此楼(FDA 官方照片,公共领域)。The FDA's Building 66 at its White Oak campus in Silver Spring, Maryland, home of the Center for Devices and Radiological Health (CDRH). Photo: U.S. Food and Drug Administration, public domain.
GSM Global · 本篇英文版 English edition of this story